
Variant Creutzfeldt-Jakob Disease (vCJD), a devastating neurodegenerative disorder, stands as a stark reminder of the intricate and often perilous relationship between humans and the food chain. Unlike its sporadic counterpart, vCJD emerged as a new variant, its genesis undeniably linked to the consumption of meat contaminated with the infectious agent responsible for bovine spongiform encephalopathy (BSE), commonly known as “mad cow disease.” This article will delve into the complex web of factors that contribute to the occurrence of vCJD, offering a factual exploration akin to consulting a comprehensive encyclopedia.
At the heart of vCJD lies a misfolded protein, a rogue agent known as a prion. This isn’t a virus or a bacterium, but a normal protein found in the brain that has adopted an aberrant, infectious shape. Think of it like a key that no longer fits its lock, but instead, upon encountering the correct lock, forces it to change its shape to match the rogue key. This misfolding is the primal wound, the spark that ignites the cascade of neurological damage.
What is a Prion?
Prions are misfolded isoforms of the cellular prion protein (PrPC). In its normal form, PrPC is a soluble protein involved in various cellular functions, although its precise role remains an area of active research. The abnormal, infectious form is designated as PrPSc (for scrapie, a related prion disease in sheep). PrPSc possesses a distinct three-dimensional structure, characterized by an increased beta-sheet content. This structural difference renders it resistant to degradation by proteases, the enzymes that typically break down cellular proteins.
The Mechanism of Prion Replication: A Chain Reaction
The insidious nature of prions lies in their ability to induce the misfolding of normal PrPC molecules into the infectious PrPSc form. This is not a process of genetic replication, as seen with viruses or bacteria. Instead, it’s a conformational conversion, a domino effect where a single PrPSc molecule acts as a template to convert neighboring PrPC molecules. This autocatalytic process, once initiated, leads to an exponential accumulation of PrPSc.
- Conformational Change: The key to prion infectivity is the change in protein shape. PrPC typically contains alpha-helical structures, while PrPSc is rich in beta-pleated sheets. This fundamental structural shift confers the protein’s resistance to degradation and its templating ability.
- Aggregations and Fibril Formation: As PrPSc accumulates, it begins to aggregate, forming insoluble structures known as amyloid fibrils. These fibrils are a hallmark of prion diseases and are believed to contribute to cellular dysfunction and neuron death.
- Synaptic Dysfunction: The accumulation of PrPSc can disrupt the normal function of synapses, the junctions between neurons where communication occurs. This disruption is an early sign of the neurological damage that characterizes vCJD.
Prion Strain Diversity: A Subtle Menace
While the basic mechanism of prion replication is consistent, there exists a phenomenon known as prion strain diversity. Different strains can arise from subtle variations in the conformation of PrPSc. These variations can influence the incubation period of the disease, the specific brain regions affected, and the clinical signs observed. In the context of vCJD, the prion strain is believed to be derived from the one responsible for BSE, but its passage through cattle and then into humans may have subtly altered its characteristics.
Recent research has shed light on the causes of variant Creutzfeldt-Jakob disease (vCJD), a rare and fatal neurodegenerative disorder linked to the consumption of beef infected with bovine spongiform encephalopathy (BSE), commonly known as mad cow disease. For a deeper understanding of the implications of this disease and its connection to prion infections, you can read a related article that explores the latest findings and preventive measures. Check it out here: Freaky Science.
The BSE-vCJD Link: A Culinary Catastrophe
The emergence of vCJD in the early 1990s was a watershed moment in our understanding of zoonotic prion diseases. The scientific community rapidly established a strong link between the rise in BSE in cattle and the appearance of this new human illness. This connection painted a grim picture of how dietary practices could inadvertently bridge the species barrier, exposing humans to a novel and deadly pathogen.
The Genesis of BSE in Cattle: Industrial Farming and Feed
The roots of the BSE epidemic can be traced back to changes in cattle farming practices in the United Kingdom during the latter half of the 20th century. The widespread inclusion of rendered animal protein, specifically meat-and-bone meal, in cattle feed was a key culprit. Historically, ruminants were primarily herbivores. However, the economics of intensive farming led to the practice of feeding cattle processed byproducts from other animals, including sheep and cattle.
- Recycling of Prions: The rendering process, intended to sterilize and process animal carcasses, proved insufficient to inactivate prions. If infected animal material was included in the rendering process, the prions could survive and thus be reintroduced into the food chain.
- Scrapie Contribution: It is hypothesized that prions from scrapie-infected sheep were inadvertently incorporated into the animal feed, thus initiating the BSE epidemic in cattle. The species barrier, generally considered a protective mechanism, was overcome in this instance.
- The Bovine-Specific Prion Strain: Over time, the prion strain within the cattle population evolved, adapting to its bovine host to become the distinct BSE agent. This adaptation, though not fully understood, highlights the potential for prion strains to mutate or adapt when transmitted to a new host species.
Transmission to Humans: A Dietary Betrayal
The primary route of human exposure to the BSE agent leading to vCJD is believed to have been through the consumption of beef products contaminated with PrPSc. The practice of feeding cattle meat-and-bone meal that contained infected cattle tissue meant that the infectious agent was cycling within the herd and subsequently entering the human food supply.
- Consumption of Infected Tissues: The brain, spinal cord, and certain other tissues of cattle infected with BSE are known to have the highest concentration of the infectious prion. Consumption of these tissues, particularly in processed foods where their origin might be obscured, posed a significant risk.
- The Incubation Period: A Silent Threat: A terrifying aspect of prion diseases is their incredibly long incubation periods. This means that individuals could have been exposed to the BSE agent years, even decades, before the onset of symptoms. This protracted period of asymptomatic infection made it incredibly challenging to track the extent of the outbreak and implement effective control measures.
- Mechanisms of Entry: While ingestion is considered the primary route, the possibility of other, less common routes of transmission cannot be entirely ruled out. However, the overwhelming evidence points towards dietary exposure as the main driver of the vCJD epidemic.
Susceptibility and Genetic Predisposition: The Human Factor
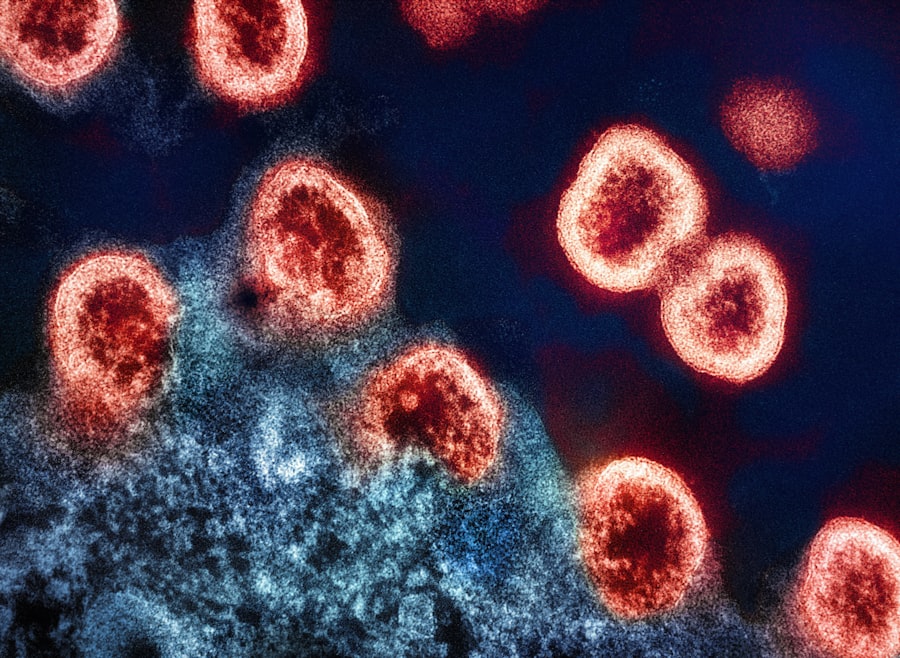
While the exposure to the BSE agent is the trigger for vCJD, not every individual exposed develops the disease. This suggests that a combination of factors, including genetic predisposition and the specific dose and form of the infectious prion encountered, play a crucial role in determining susceptibility.
The Role of the PRNP Gene: A Genetic Blueprint
The human gene responsible for producing the cellular prion protein is known as the prion protein gene (PRNP). Polymorphisms, variations in the DNA sequence of this gene, can influence an individual’s susceptibility to prion diseases.
- Methionine/Valine Polymorphism at Codon 129: The most well-studied polymorphism in the PRNP gene occurs at codon 129, which dictates the amino acid at position 129 of the prion protein. Individuals can be homozygous for methionine (Met/Met), homozygous for valine (Val/Val), or heterozygous (Met/Val).
- Homozygosity for Methionine: Research has consistently shown that individuals who are homozygous for methionine at codon 129 of the PRNP gene are significantly more susceptible to developing vCJD. This genetic makeup appears to facilitate the conversion of normal PrPC to the infectious PrPSc.
- Heterozygosity and Reduced Risk: Individuals who are heterozygous (Met/Val) appear to have a lower susceptibility, although they are not entirely immune.
- Homozygosity for Valine: Individuals homozygous for valine (Val/Val) are considered to have the lowest risk of developing vCJD.
The Influence of Prion Strain and Dose: A Complex Equation
Beyond individual genetics, the characteristics of the infecting prion also influence disease development. The specific strain of the BSE agent and the amount of infectious material ingested are likely to be critical determinants of whether an individual develops vCJD.
- Endogenous Prion Conversion: The exact mechanism by which ingested PrPSc initiates the disease process in humans is still being elucidated. It is hypothesized that swallowed PrPSc enters the lymphoreticular system and then travels to the brain, where it encounters endogenous PrPC and triggers the conformational conversion.
- Dose-Dependent Exposure: A higher dose of the BSE agent, encountered through more frequent or larger consumption of contaminated products, would logically increase the probability of infection and subsequent disease development.
Global Public Health Responses and Eradication Efforts: A Battle Against the Invisible
The recognition of vCJD as a human health crisis spurred significant and coordinated public health interventions worldwide. The focus shifted from merely treating the disease, for which no cure exists, to preventing its further spread and ultimately eradicating the conditions that allowed its emergence.
Stricter Regulations on Animal Feed: Closing the Loophole
One of the most critical steps in combating vCJD was the implementation of stringent regulations regarding animal feed. The goal was to sever the cycle of prion transmission within the agricultural sector.
- Bans on Meat-and-Bone Meal: Most industrialized nations implemented bans on the use of meat-and-bone meal derived from ruminants in the feed of other ruminants. This effectively removed the primary route by which prions were recycled within cattle populations.
- Specified Risk Materials (SRMs) Removal: Regulations were also established to remove “specified risk materials” (SRMs) from the food chain. These are tissues that are known to harbor high concentrations of prions, such as the brain, spinal cord, and tonsils. This measure aimed to reduce the likelihood of accidental human exposure to infectious agents present in meat products.
- Enhanced Surveillance and Testing: Improved surveillance systems were put in place to monitor cattle populations for signs of BSE. This included rigorous testing of cattle, particularly those showing neurological signs, to detect the presence of the BSE agent.
Controls on Food Production and Distribution: Safeguarding the Supply
Beyond feed regulations, measures were also implemented to ensure the safety of the wider food production and distribution network.
- Traceability and Identification Systems: Robust systems for the identification and traceability of cattle were introduced. This allows authorities to track the origin of meat products and recall potentially contaminated items if an outbreak is detected.
- Public Education Campaigns: Public education campaigns were crucial in informing consumers about the risks associated with BSE and vCJD and the measures being taken to ensure food safety. These campaigns aimed to rebuild public confidence in the food supply.
- International Cooperation: The global nature of the food trade necessitated international cooperation and harmonization of regulations. Organizations like the World Organisation for Animal Health (OIE) played a vital role in setting standards and facilitating information exchange.
Recent studies have shed light on the causes of variant Creutzfeldt-Jakob disease, a rare and fatal neurodegenerative condition linked to the consumption of contaminated beef products. For a deeper understanding of the mechanisms behind this disease and its connection to prion proteins, you can explore a related article that discusses the implications of these findings in more detail. This insightful piece can be found at Freaky Science, where it delves into the ongoing research and potential preventive measures.
Ongoing Research and Future Prospects: A Vigilant Outlook
| Aspect | Details |
|---|---|
| Disease Name | Variant Creutzfeldt-Jakob Disease (vCJD) |
| Cause | Infection by abnormal prion proteins (PrP^Sc) |
| Primary Source | Consumption of Bovine Spongiform Encephalopathy (BSE) contaminated beef |
| Transmission Mode | Oral ingestion of infected tissue; rare cases via blood transfusion |
| Incubation Period | Typically 10-15 years, can vary widely |
| Pathology | Spongiform degeneration of brain tissue, accumulation of prions |
| Symptoms | Psychiatric symptoms, sensory disturbances, ataxia, dementia |
| Diagnosis | Clinical assessment, MRI, tonsil biopsy, detection of prions in tissue |
| Prevention | Control of BSE in cattle, blood donor screening, avoiding high-risk tissues |
| Prognosis | Fatal within months to a few years after symptom onset |
Despite significant progress in controlling vCJD, the threat of prion diseases, including potential novel variants, remains a concern. Continuous research is essential to deepen our understanding of these complex disorders and to develop effective countermeasures.
Understanding Prion Biology: Unraveling the Mysteries
Further research into the fundamental biology of prions is crucial for developing new diagnostic and therapeutic strategies.
- Structural Dynamics of Prion Misfolding: Detailed investigation into the precise molecular mechanisms by which PrPC misfolds into PrPSc is ongoing. This could unlock potential targets for drugs that could inhibit or reverse this process.
- Prion Strain Characterization: Continued research into the diversity of prion strains and the factors that influence their emergence and transmissibility is vital. This knowledge can help predict and prepare for potential new threats.
- The Role of Co-factors: Scientists are exploring the role of other cellular molecules that might act as co-factors in prion replication. Identifying these co-factors could provide additional avenues for therapeutic intervention.
Developing Diagnostics and Therapeutics: The Quest for Solutions
The development of effective diagnostic tools and therapeutic interventions for prion diseases is a paramount goal.
- Early Detection Methods: Current diagnostic methods for vCJD, such as brain imaging and tonsil biopsy, are often only effective once the disease has progressed significantly. Research is focused on developing more sensitive and less invasive diagnostic tests that can detect prions or their effects early in the incubation period. This could involve identifying biomarkers in blood or cerebrospinal fluid.
- Therapeutic Strategies: The absence of a cure for vCJD is a profound challenge. Therapeutic research is exploring various approaches, including:
- Anti-prion drugs: Compounds designed to inhibit the conversion of PrPC to PrPSc or to promote the clearance of PrPSc.
- Immunotherapies: Strategies aimed at stimulating the immune system to clear prions.
- Gene therapy: Exploring ways to modulate the expression of the PRNP gene or to introduce protective genetic variations.
Vigilance Against Emerging Threats: A Guarded Future
The possibility of new prion diseases emerging, either through further adaptation of existing agents or through novel zoonotic transmissions, necessitates a continued state of vigilance.
- Monitoring Animal Health: Ongoing surveillance of animal populations for the presence of known and emerging prion diseases remains essential. This includes monitoring wildlife populations that could act as reservoirs for prion agents.
- Cross-Species Transmission Studies: Understanding the potential for cross-species transmission of prions is critical. Research involving controlled experimental transmissions can help assess the risk of novel agents jumping to humans.
- Preparedness for Future Outbreaks: Maintaining robust public health infrastructure and research capabilities is crucial for responding effectively to any future prion disease outbreaks. This involves preparedness for rapid detection, containment, and public communication.
In conclusion, exploring the causes of variant Creutzfeldt-Jakob disease is a journey through the complex interplay of biology, agriculture, and public health. From the molecular dance of misfolded proteins to the global efforts to safeguard our food supply, understanding vCJD offers profound lessons about our interconnectedness with the natural world and the importance of continuous scientific inquiry and vigilant public health practices.
FAQs
What is variant Creutzfeldt-Jakob disease (vCJD)?
Variant Creutzfeldt-Jakob disease (vCJD) is a rare, degenerative, and fatal brain disorder caused by abnormal prion proteins. It is a form of Creutzfeldt-Jakob disease linked to the consumption of contaminated beef products infected with bovine spongiform encephalopathy (BSE), also known as “mad cow disease.”
What causes variant Creutzfeldt-Jakob disease?
vCJD is caused by the ingestion of prion-infected beef products. The abnormal prions cause damage to brain tissue, leading to neurological symptoms and ultimately death. The disease is not caused by bacteria or viruses but by misfolded proteins that induce other proteins to misfold.
How is variant Creutzfeldt-Jakob disease transmitted?
The primary route of transmission is through eating beef products contaminated with BSE prions. There is no evidence that vCJD spreads through casual contact, respiratory droplets, or blood transfusions in most cases, although rare instances of transmission via blood transfusion have been documented.
What are the symptoms of variant Creutzfeldt-Jakob disease?
Symptoms typically include psychiatric problems such as depression and anxiety, sensory disturbances, coordination difficulties, memory loss, and cognitive decline. As the disease progresses, patients experience severe neurological impairment, leading to coma and death.
Is there a cure or treatment for variant Creutzfeldt-Jakob disease?
Currently, there is no cure or effective treatment for vCJD. Medical care focuses on alleviating symptoms and providing supportive care. Research continues to explore potential therapies, but the disease remains invariably fatal.